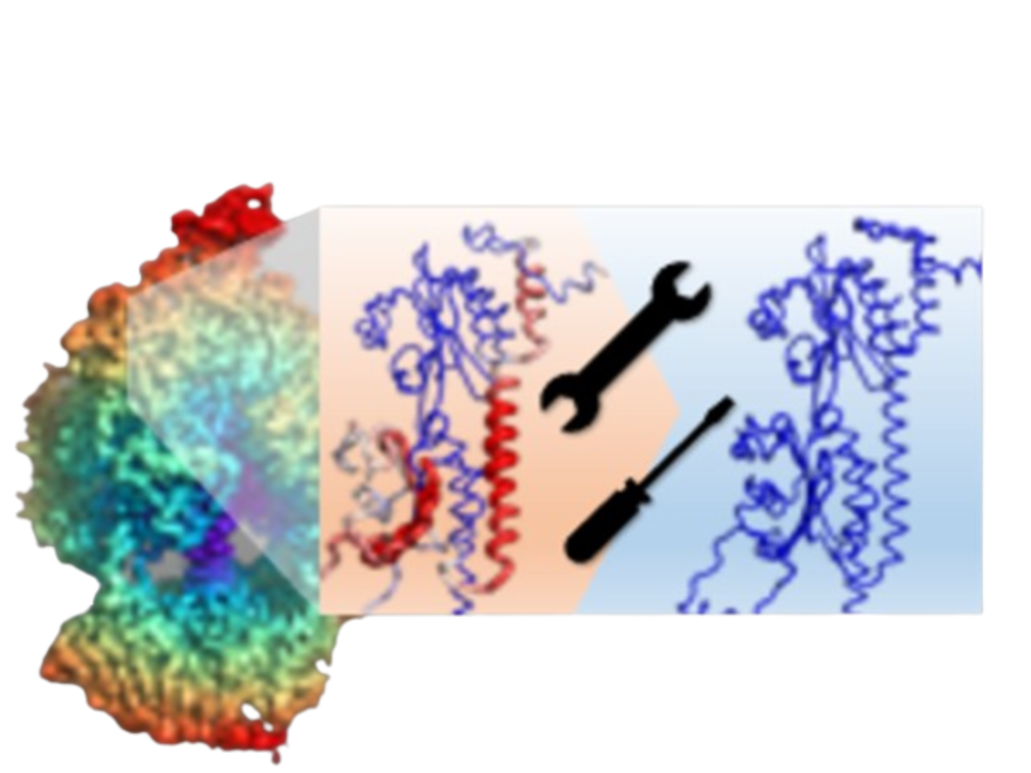
DAQ-refine is a protocol using DAQ score to evaluate protein models from cryo-EM maps and employs a modified AlphaFold2 to refine regions with potential errors.
If encounter problems, please contact Daisuke Kihara (dkihara@purdue.edu), Genki Terashi(gterashi@purdue.edu) or Xiao Wang (xiaowang20140001@gmail.com)
Example EMD-0837 chain C Input Map file:emd_0837.map.gz
Input protein file:6l54C.pdb
Input sequence:6l54C.fasta
Result Example:Result Example
Example EMD-31563 chain B Input Map file:emd_31563.map.gz
Input protein file:7fetB.pdb
Input sequence:7fetB.fasta
Result Example:Result Example
Tutorial
References:
Terashi, G., Wang, X., Kihara, D.. Protein model refinement for cryo-EM maps using AlphaFold2 and the DAQ score. Acta Cryst. D79 (2022). https://doi.org/10.1107/S2059798322011676
Terashi, G., Wang, X., Maddhuri Venkata Subramaniya, S.R., Kihara, D.. Residue-wise local quality estimation for protein models from cryo-EM maps. Nat Methods 19.9 (2022). https://doi.org/10.1038/s41592-022-01574-4
Please simply click "Schedule Job" when you filled all input fields.
Please see the instruction for How to Preparing the PDB file.
Please see the instruction for How to Preparing the fasta file.
Due to limited computational resources, we can only process proteins with a maximum length of 1500 amino acids for each chain. Also, the processing time can be long, please be patient and wait for the result.
If your turn in a job with the structure >20 chains, it could take one week or more time to finish.
If the chain in the fasta file contains 'X' only, we would not process this chain and delete it in the final result.
Please select the alignment strategy for PDB file and fasta sequence. We strongly recommend you align the 2 files for chains mannually(keep the same order for chains in 2 files), a simple tutorial here. If you are not sure, please select "Smith Waterman alignment".
0/500
This note will help you distinguish jobs later.